

SECTION 1
Historical Context & Motivation
The relationship between electrical work and chemical change was not always obvious. In the late eighteenth century, scientists began to realize that certain chemical reactions could produce a sustained electric current, while externally applied electrical energy could drive otherwise non-spontaneous reactions. This dual observation raised a profound question: is there a single thermodynamic quantity that governs both the direction and the magnitude of electrochemical processes? The answer came through a convergence of experimental ingenuity and theoretical insight spanning more than a century, ultimately linking the measurable voltage of an electrochemical cell to the Gibbs free energy of the underlying reaction.
1800 • Volta's Pile
1834 • Faraday's Laws of Electrolysis
1876 • Gibbs Free Energy Formulated
1889 • The Nernst Equation
1923 • Debye–Hückel Theory
These advances converged on a central insight: the voltage produced by an electrochemical cell is not merely an electrical curiosity but a direct, quantitative window into the thermodynamic driving force of a reaction. The question that motivates this lesson is straightforward yet powerful—how does the cell potential E° connect to the Gibbs free energy change ΔG°, and what does that connection tell us about spontaneity, equilibrium, and the maximum useful work a reaction can perform?
SECTION 2
Core Principles & Definitions
Before diving into the mathematics, it is essential to establish the foundational definitions that underpin electrochemistry and its thermodynamic interpretation. An electrochemical cell consists of two half-cells, each containing an electrode immersed in an electrolyte; electrons flow through an external circuit from the anode (oxidation) to the cathode (reduction), while ions migrate through a salt bridge or porous barrier to maintain electrical neutrality. The standard cell potential (E°cell) is the electromotive force measured when all species are at standard-state conditions—1 M concentrations for solutes, 1 atm partial pressures for gases, and pure solids or liquids at 25 °C.
Cell Potential (E°cell)
Gibbs Free Energy (ΔG°)
Faraday's Constant (F)
Moles of Electrons (n)
Equilibrium Constant (K)
KEY TAKEAWAY
SECTION 3
Visual Explanation: The Galvanic Cell
The diagram above illustrates the classic Daniell cell, which serves as a prototype for understanding electrochemical spontaneity. Zinc is oxidized at the anode because it has a more negative standard reduction potential (E° = −0.76 V) compared to copper (E° = +0.34 V). The difference, E°cell = +0.34 − (−0.76) = +1.10 V, is positive, confirming that the reaction is spontaneous as written. Every positive volt of cell potential corresponds to a negative ΔG°, and therefore to a reaction that can do useful electrical work on its surroundings. The salt bridge is essential: without it, charge would build up in each half-cell, quickly halting the reaction. By allowing anions to migrate toward the anode compartment and cations toward the cathode compartment, the salt bridge maintains electrical neutrality and permits continuous current flow.
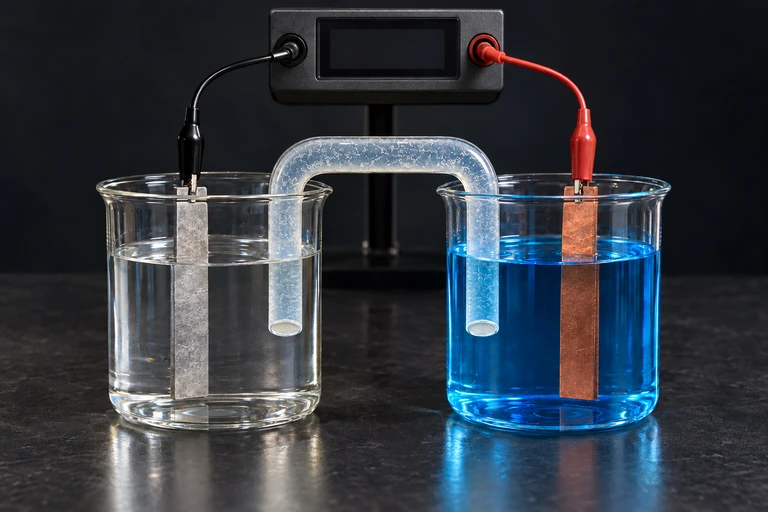 A zinc–copper cell connects two half-cells through an external circuit and a salt bridge.
SECTION 4
Mathematical Framework
The relationship between cell potential and Gibbs free energy is derived by recognizing that the maximum non-expansion work an electrochemical cell can perform is equal to the charge transferred multiplied by the potential difference driving that transfer. When n moles of electrons pass through a potential difference E, the electrical work is welec = nFE. Because the Gibbs free energy change equals the maximum non-expansion work at constant T and P (with a sign convention that ΔG is negative for spontaneous work-producing processes), we arrive at the fundamental equation.
GIBBS FREE ENERGY–CELL POTENTIAL RELATIONSHIP
ΔG° = standard Gibbs free energy change (J mol⁻¹); n = moles of electrons transferred; F = Faraday's constant (96 485 C mol⁻¹); E°cell = standard cell potential (V). Note: 1 V × 1 C = 1 J.
The negative sign ensures internal consistency: a positive E°cell (spontaneous galvanic cell) yields a negative ΔG° (thermodynamically favorable), while a negative E°cell implies a positive ΔG° (non-spontaneous; the cell operates only if external energy is supplied, as in electrolysis).
CONNECTION TO THE EQUILIBRIUM CONSTANT
R = 8.314 J mol⁻¹ K⁻¹; T = temperature in Kelvin; K = thermodynamic equilibrium constant. Combining with ΔG° = −nFE° gives: E° = (RT / nF) ln K.
NERNST EQUATION (NON-STANDARD CONDITIONS)
E = cell potential under actual conditions; Q = reaction quotient. At 25 °C, this simplifies to E = E° − (0.0257 V / n) ln Q, or equivalently E = E° − (0.0592 V / n) log Q.
COMBINED RELATIONSHIP AT 25 °C
Derived by setting ΔG° = −nFE° = −RT ln K and solving for E°. This equation directly links the standard cell potential to the equilibrium constant without needing ΔG° explicitly.
Sign Convention Check
SECTION 5
The Thermodynamic Triangle: E°, ΔG°, and K
One of the most powerful conceptual tools in AP Chemistry is the thermodynamic triangle connecting three quantities: the standard cell potential E°cell, the standard Gibbs free energy change ΔG°, and the equilibrium constant K. Knowing any one of these three values allows you to calculate the other two, because they are all manifestations of the same thermodynamic driving force. The diagram below maps these interconversions and highlights the sign relationships that govern spontaneity.
| Condition | E°cell | ΔG° | K | Interpretation |
|---|---|---|---|---|
| Spontaneous | > 0 | < 0 | > 1 | Products favored; galvanic cell produces work |
| At equilibrium | = 0 | = 0 | = 1 | No net reaction; dead battery |
| Non-spontaneous | < 0 | > 0 | < 1 | Reactants favored; requires external energy (electrolysis) |
Summary of sign relationships among the three thermodynamic quantities
SECTION 6
Worked Example
Consider a galvanic cell based on the following reaction: 2 Ag⁺(aq) + Fe(s) → 2 Ag(s) + Fe²⁺(aq). Given the standard reduction potentials E°(Ag⁺/Ag) = +0.80 V and E°(Fe²⁺/Fe) = −0.44 V, calculate E°cell, ΔG°, and K at 25 °C.
Calculating E°, ΔG°, and K for a Silver–Iron Cell
- 1
Step 1 — Identify the half-reactions and assign anode/cathode
Ag⁺ has the more positive standard reduction potential (+0.80 V), so Ag⁺ is reduced at the cathode. Fe has the more negative reduction potential (−0.44 V), so Fe is oxidized at the anode. The half-reactions are: Cathode: 2 Ag⁺(aq) + 2 e⁻ → 2 Ag(s); Anode: Fe(s) → Fe²⁺(aq) + 2 e⁻.n = 2 mol e⁻ transferred - 2
Step 2 — Calculate the standard cell potential
E°cell = E°cathode − E°anode = (+0.80 V) − (−0.44 V) = +1.24 V. The positive value confirms the reaction is spontaneous under standard conditions.E°cell = +1.24 V - 3
Step 3 — Calculate ΔG°
ΔG° = −nFE° = −(2 mol)(96 485 C mol⁻¹)(+1.24 V) = −(2)(96 485)(1.24) J = −239 283 J ≈ −239.3 kJ. The large negative value indicates that this reaction releases a substantial amount of free energy.ΔG° ≈ −239 kJ - 4
Step 4 — Calculate K
Using ΔG° = −RT ln K, rearrange to ln K = −ΔG° / RT = −(−239 283 J) / [(8.314 J mol⁻¹ K⁻¹)(298 K)] = 239 283 / 2477.6 ≈ 96.6. Therefore K = e96.6 ≈ 1.3 × 1041. This enormous K value confirms that the reaction goes essentially to completion.K ≈ 1.3 × 10⁴¹ - 5
Step 5 — Verify consistency
Check: E° > 0 ✓ → ΔG° < 0 ✓ → K >> 1 ✓. All three quantities are consistent with a strongly product-favored, spontaneous reaction. This internal consistency check is a valuable exam strategy.All signs consistent ✓
SECTION 7
Strengths, Limitations, and Common Misconceptions
While the relationship ΔG° = −nFE° is elegant and widely applicable, students frequently encounter pitfalls when applying it. Understanding both the power and the boundaries of this equation is essential for success on the AP exam and for developing genuine chemical intuition.
| Strength | Limitation / Misconception | Clarification |
|---|---|---|
| Directly links measurable voltage to thermodynamic spontaneity | Students sometimes think E° changes when the balanced equation is multiplied by a factor | E° is an intensive property and does not change with stoichiometric coefficients. However, ΔG° does change because n changes. |
| Provides a route to K without needing ΔH° and ΔS° separately | Assumes standard conditions; real cells operate at non-standard concentrations | Use the Nernst equation (E = E° − (RT/nF) ln Q) to account for actual conditions. |
| Connects electrochemistry to chemical equilibrium via K | A common error is confusing thermodynamic spontaneity with reaction rate | ΔG° tells you whether a reaction can occur, not how fast. Kinetics (activation energy, catalysts) governs rate. |
| Works for any redox reaction, not just simple metal-ion cells | Students may forget to balance electrons before determining n | Always write and balance both half-reactions first, then identify n from the balanced full equation. |
Key strengths, common misconceptions, and clarifications
KEY TAKEAWAY
SECTION 8
Connections to Advanced Theory
The equation ΔG° = −nFE° is a gateway to deeper topics in physical chemistry and materials science. In this section, we briefly survey how the standard-condition treatment extends to more sophisticated frameworks that you may encounter in advanced coursework or on challenging AP free-response questions.
| AP-Level Concept | Advanced Extension |
|---|---|
| ΔG° = −nFE° at 25 °C | Temperature-dependent ΔG via Gibbs-Helmholtz equation: ΔG = ΔH − TΔS; at non-standard temperatures, E° changes and can be predicted using the temperature coefficient dE°/dT = ΔS°/(nF) |
| Nernst equation with concentration-based Q | In real solutions, activities (not concentrations) should replace molarity terms. The Debye-Hückel theory provides activity coefficients for dilute ionic solutions. |
| Standard hydrogen electrode (SHE) as reference | Other reference electrodes (Ag/AgCl, saturated calomel) are used in research. Potentials must be converted to the SHE scale for thermodynamic calculations. |
| Galvanic vs. electrolytic cells | Overpotential and kinetic barriers in electrolysis mean that the actual voltage required exceeds E°cell; these are described by the Butler-Volmer and Tafel equations in electrochemical kinetics. |
AP-level concepts and their advanced extensions
For the AP exam, the most critical extension is the Nernst equation, which generalizes the standard-state relationship to any set of conditions. When a battery discharges, Q increases as products accumulate and reactants are consumed. Consequently, E decreases from E° toward zero. When E reaches zero, Q equals K, ΔG equals zero, and the cell is at equilibrium—the battery is 'dead.' This elegant progression, from a fully charged galvanic cell to a dead battery to a rechargeable electrolytic cell (when external voltage is applied), is all encoded in the interplay between ΔG, E, and Q.
AP Exam Tip
SECTION 9
Practice Problems
Question 1: CONCEPTUAL
- A. ΔG° > 0 and K > 1
- B. ΔG° < 0 and K > 1
- C. ΔG° < 0 and K < 1
- D. ΔG° > 0 and K < 1
Show answer (B)
Question 2: BASIC CALCULATION
- A. +1.10 V
- B. +1.62 V
- C. −1.62 V
- D. +2.72 V
Show answer (B)
Question 3: INTERMEDIATE
- A. −386 kJ
- B. −579 kJ
- C. −1 158 kJ
- D. −193 kJ
Show answer (C)
Question 4: APPLIED
10 ptsShow answer
Question 5: CRITICAL THINKING
10 ptsShow answer
SUMMARY
Summary
Summary
The central equation of this lesson, ΔG° = −nFE°, establishes a direct, quantitative bridge between the standard cell potential measured in volts and the Gibbs free energy change measured in joules. A positive E°cell indicates a spontaneous galvanic cell (negative ΔG°), while a negative E°cell signals a non-spontaneous process requiring external energy input. The variable n (moles of electrons) and Faraday's constant F (96 485 C mol⁻¹) serve as the conversion factors between the electrical and thermodynamic domains.
The thermodynamic triangle connecting E°, ΔG°, and K allows you to compute any one quantity from either of the other two. The Nernst equation extends this framework to non-standard conditions by incorporating the reaction quotient Q. Remember that E° is intensive (does not change when you scale the equation), while ΔG° is extensive (scales with n). Master these relationships and you will be prepared to tackle any electrochemistry–thermodynamics question on the AP Chemistry exam.